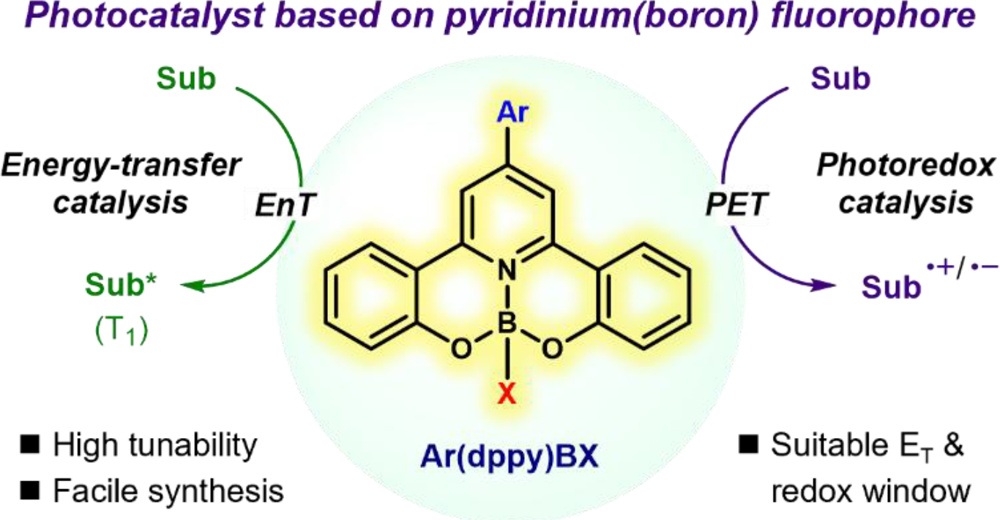
Novel photocatalyst
A modular, structurally tunable fluorophore for diverse photochemical transformations.
The Story
When I took over this project, most of the hard synthetic work was already behind it. Lutao Bai had made the catalyst, and he had already screened the redox reactions it could drive—there was a working molecule in hand and a set of transformations it was good for. What we did not have was an answer to the obvious question: what is this thing, really?
We were also honest with ourselves about the stakes. A new fluorophore that merely does what 4CzIPN does, under a different name, is a substitution rather than a contribution—and a substitution would not have carried this work very far. The catalyst had to be more than a replacement. So we made a deliberate decision early on to spend our effort in two directions instead of one: to establish what this molecule is genuinely capable of beyond simple photoredox catalysis, and to understand its mechanism at a level that no one could argue with.
The first of these was energy transfer. I worked through the photosensitization behavior of the catalyst in depth and established that it is a genuinely excellent triplet sensitizer—in the reactions where the benchmark is a precious-metal complex, ours matches the strongest iridium photosensitizers, and in several cases goes beyond them. That comparison is the point: everything the iridium complexes are good at, this molecule is also good at, while being metal-free, inexpensive, and assembled in a few straightforward steps from simple building blocks. In a field where the best triplet sensitizers are still iridium complexes, that is not a small thing to be able to say.
The second direction was the mechanism, and it is where I spent most of my time. I essentially emptied the spectroscopic toolkit onto this system: transient absorption on the nanosecond and femtosecond timescales, steady-state EPR, and our own home-built U-PSD TREPR, together with time-resolved kinetics for every species we could catch. What came out was not a single lucky observation but a coherent set of mechanistic conclusions—which excited state is actually responsible for the electron transfer, how the catalyst degrades and how to stop it, what really governs the rate of the catalytic cycle, and why the catalyst with the larger thermodynamic driving force is sometimes the worse one. I will not enumerate them all here; they are the substance of the paper.
If there is a lesson I keep from this project, it is about ambition. The catalyst arrived on my bench as a finished molecule. The work was deciding that it deserved to be more than that.
Academic Summary
This work establishes the pyridinium(boron) motif—specifically the modular Ar(dppy)BX scaffold—as a broadly tunable, metal-free organic photocatalyst, translating our group’s long-standing N-boryl pyridyl anion chemistry from a ground-state reagent into a covalently preorganized light-harvesting platform.1 The design addresses a concrete limitation in the field: while benchmark fluorophores such as 4CzIPN combine excellent excited-state lifetimes with wide redox windows, their donor–acceptor architecture offers little room for independent tuning—modulating the redox potential generally requires altering multiple donor units while the acceptor remains fixed. The Ar(dppy)BX framework circumvents this constraint by exposing two orthogonal handles, the aryl group (Ar) on the pyridinium core and the substituent (X) at the boron center, which permit the excited-state reduction and oxidation potentials to be adjusted separately.
The model catalyst Ph(dppy)BOMe (1a) was prepared in four steps (41% overall) and structurally authenticated by single-crystal XRD. Cyclic voltammetry established a reversible reduction at E₁/₂(1a/1a•⁻) = −1.53 V and an irreversible oxidation at Eₚ,ₐ(1a•⁺/1a) = +1.57 V vs SCE; UV–vis, fluorescence, and phosphorescence spectroscopy located the absorption maximum at 385 nm with emission at 466 nm and phosphorescence at 477/502 nm, giving E_S ≈ 2.93 eV and E_T ≈ 2.71 eV (62 kcal/mol). Application of the Rehm–Weller treatment yielded E_red* = +1.39 V and E_ox* = −1.35 V vs SCE. Time-correlated single-photon counting measured a singlet lifetime of 4.1 ns, while nanosecond transient absorption revealed a longer-lived triplet species (λ_max = 528 nm, τ = 6.1 μs) that was quenched under aerated conditions. TD-DFT hole–electron analysis showed electron transfer from the phenolate to the pyridine subunit upon S₀ → S₁ excitation, with spin density delocalized across both fragments in T₁—a partial charge separation that rationalizes the molecule’s dual photoredox and photosensitization character. Relative to 4CzIPN, 1a offers a wider redox window in both quenching directions and a higher triplet energy, at the cost of shorter excited-state lifetimes that nonetheless remain fully competent for bimolecular chemistry.
The photochemical behavior of 1a was then dissected with substrate probes rather than inferred from thermodynamics alone. Reductive quenching by pempidine proceeded with k_Q = 7.8 × 10⁹ M⁻¹ s⁻¹; steady-state EPR detected the resulting radical anion 1a•⁻ and confirmed its identity against an independently prepared sample generated by chemical reduction with KC₈. The transient counter-radical, the pempidine radical cation, is invisible to conventional EPR and was captured instead by our home-built U-PSD TREPR, which resolved its hyperfine pattern (a_N = 2.04 mT, 3a_H = 2.75 mT) and a lifetime of approximately 7 μs. The absorption–emission spin polarization of the signal reported the formation of a singlet radical pair, establishing that the S₁ state—not T₁—governs the electron transfer. In the oxidative direction, quenching by methyl viologen (k_q = 1.2 × 10¹⁰ M⁻¹ s⁻¹) produced MV•⁺, whose persistence in transient absorption allowed the competing charge-recombination (k_CR = 8.0 × 10⁹ M⁻¹ s⁻¹) and catalyst-degradation (~15% of 1a•⁺ per excitation pulse) pathways to be quantified separately. Finally, trans-stilbene quenching (k_Q = 8.64 × 10⁹ M⁻¹ s⁻¹) left the fluorescence lifetime untouched, confirming that 1a operates as a genuine triplet sensitizer.
These capabilities translated directly into synthetic performance. In direct SET oxidations—the decarboxylative alkylation of a proline derivative and the formal [3 + 2] cycloaddition of tBu₃P with terminal alkynes—1a matched 4CzIPN. In reductive quenching cycles it surpassed it, delivering the defluorinative alkylation product of benzotrifluoride in 83% yield against 19% for 4CzIPN and, notably, without requiring the thiophenol HAT co-catalyst, while a Birch-type reduction of anthracene also proceeded more efficiently. The advantage was most pronounced in energy-transfer catalysis: with Murakami’s probe molecule, 1a funneled the reaction almost exclusively to the EnT product (15b, 86%) rather than the SET product (15a, 3%), and its 62 kcal/mol triplet energy allowed an intramolecular [2 + 2] cycloaddition of a high-lying substrate to proceed with efficiency comparable to [Ir(dF-CF₃-ppy)₂(dtbbpy)]PF₆. Three transformations that conventionally demand iridium or ultraviolet light—the Rovis oxime photoisomerization, the [2 + 2] cycloaddition of 4-hydroxycoumarins, and the Huang ring expansion of oximes—were driven to completion by 1a under 400 nm irradiation, whereas 4CzIPN failed entirely.
A further demonstration of the platform’s utility is the photocatalyzed switchable 1e/2e oxidation of amines, in which a single catalyst and a single substrate class are diverted toward divergent products by changing the base and the irradiation wavelength. With Cs₂CO₃ and 450 nm light, the reaction furnishes α-arylation products (1e pathway) across cyclic and acyclic anilines and a range of dicyanobenzene coupling partners, with regioselectivity for the less hindered α-position; switching to NaOAc and 400 nm reverses the chemoselectivity to α-cyanation (2e pathway). In both directions the ratio of the targeted product to its alternative exceeds 3:1. Because the 1e/2e branch point had previously been controllable only under electrochemical conditions, this represents the first photocatalytic solution to a long-standing selectivity problem in amine functionalization.
The modularity of the scaffold was then used as a design tool rather than a curiosity. In an oxidation-initiated C–H arylation of dihydrobenzofuran, appending electron-withdrawing groups to the pyridinium core (1c, 1d) progressively raised E_red* and improved the yield from 6% (electron-rich 1e) to 65% (1d), outperforming 4CzIPN—a clean correlation between excited-state oxidizing power and catalytic outcome. The complementary series, bearing amine donors at boron (1f–h), behaved counterintuitively: although these modifications shifted E_ox* to more negative values, they also quenched fluorescence by opening an intramolecular charge-transfer state, and the catalyst with the smaller thermodynamic driving force (1h, 65%) outperformed the one with the larger (1g, 48%). Femtosecond transient absorption on a soluble surrogate resolved this apparent contradiction: the charge-transfer state decays nonradiatively on a sub-100 ps timescale (k_nr = 7.4 × 10¹⁰ s⁻¹), yet intermolecular electron transfer to methyl viologen remains efficient, and kinetic analysis showed less than 3% catalyst decomposition per cycle—a fivefold improvement over 1a—together with a slower charge recombination for 1h•⁺ (7.2 × 10⁹ M⁻¹ s⁻¹) than for 1g•⁺ (2.4 × 10¹⁰ M⁻¹ s⁻¹). The lesson is quantitative and general: in oxidative quenching cycles, catalytic throughput is governed as much by resistance to degradation and by charge-recombination kinetics as by the excited-state redox potential itself.
Collectively, this study converts a legacy organocatalytic motif into a broad-spectrum photocatalyst, and demonstrates that a compact, modular fluorophore with independently addressable donor and acceptor positions can complement—and in several challenging energy-transfer and reductive-quenching regimes, outperform—both 4CzIPN and established iridium photosensitizers.